Showing posts with label Spontaneity. Show all posts

Friday, April 23, 2021

Chapter 7.8 - Relation Between Equilibrium Constant and Gibbs Energy

In the previous section, we saw the applications of equilibrium constant. In this section, we will see how equilibrium constant is related to Gibbs energy

• In the previous chapter, we saw that a reaction will occur spontaneously in the forward direction if ΔG is negative (Details here)
• In the previous section of the present chapter, we saw that, if Qc is less than Kc, the reaction will occur spontaneously in the forward direction
◼ So there must be a relation between the three quantities:
(i) ΔG (ii) Kc (or Kp) (iii) Qc (or Qp)


Let us recall the three points that we have learnt about ΔG in the previous chapter:
(i) If ΔG is negative, the reaction is spontaneous and proceeds in the forward direction
(ii) If ΔG is positive, the reaction is non-spontaneous
    ♦ But then, the backward reaction will have a negative ΔG value
    ♦ So the backward reaction will proceed spontaneously
(iii) If ΔG is zero, the reaction is at equilibrium. There is no free energy available to drive the reaction in any direction
◼ Scientists have derived the following equation which gives the relation between ΔG, K and Q
Eq.7.4: $\mathbf\small{\rm{\Delta G= \Delta G^\circleddash + RT\,lnQ}}$
    ♦ Where: $\mathbf\small{\rm{\Delta G^\circleddash}}$ is the standard Gibbs energy

• For convenience of our present discussion,
    ♦ We write Kc (or Kp) simply as K
    ♦ We write Qc (or Qp) simply as Q

We can write an analysis of Eq.7.4 in 4 steps:
1. At equilibrium, we have: ΔG = 0
    ♦ At equilibrium, we also have Q = K
• So Eq.7.4 becomes: $\mathbf\small{\rm{0= \Delta G^\circleddash + RT\,lnK}}$
2. From this, we get:
Eq.7.5: $\mathbf\small{\rm{\Delta G^\circleddash = -RT\,lnK}}$
• Thus we get:
Eq.7.6: $\mathbf\small{\rm{lnK=\frac{-\Delta G^\circleddash}{RT}}}$
3. Taking antilog of both sides, we get:
Eq.7.7: $\mathbf\small{\rm{K=e^{\frac{-\Delta G^\circleddash}{RT}}}}$
4. Eq.7.7 gives us a relation between K and $\mathbf\small{\rm{\Delta G^\circleddash}}$
• That means, we can predict the value of K using the value of $\mathbf\small{\rm{\Delta G^\circleddash}}$
• This can be further explained in two steps:
(i) If $\mathbf\small{\rm{\Delta G^\circleddash}}$ is negative, then $\mathbf\small{\rm{\frac{-\Delta G^\circleddash}{RT}}}$ will be positive
    ♦ Consequently, $\mathbf\small{\rm{e^{\frac{-\Delta G^\circleddash}{RT}}}}$ will be greater than 1
    ♦ Consequently, K will be greater than 1
• When K is greater than 1, it means that:
    ♦ the numerator in $\mathbf\small{\rm{\frac{[C]^c[D]^d}{[A]^a[B]^b}}}$
    ♦ is greater than
    ♦ the denominator
• That means:
    ♦ concentrations of products
    ♦ will be greater than 
    ♦ concentrations of reactants
• That means:
The reaction will proceed spontaneously in the forward direction

(ii) If $\mathbf\small{\rm{\Delta G^\circleddash}}$ is positive, then $\mathbf\small{\rm{\frac{-\Delta G^\circleddash}{RT}}}$ will be negative
    ♦ Consequently, $\mathbf\small{\rm{e^{\frac{-\Delta G^\circleddash}{RT}}}}$ will be less than 1
    ♦ Consequently, K will be less than 1
• When K is less than, it means that:
    ♦ the numerator in $\mathbf\small{\rm{\frac{[C]^c[D]^d}{[A]^a[B]^b}}}$
    ♦ is less than
    ♦ the denominator
• That means:
    ♦ concentrations of products
    ♦ will be less than 
    ♦ concentrations of reactants
• That means:
The reaction will proceed spontaneously in the reverse direction

Let us see some solved examples:
Solved example 7.31
The value of ∆G⊝ for the phosphorylation of glucose in glycolysis is 13.8 kJ mol-1.
Find the value of Kc at 298 K.
Solution:
1. We have Eq.7.6: $\mathbf\small{\rm{lnK=\frac{-\Delta G^\circleddash}{RT}}}$
• Substituting the known values, we get: $\mathbf\small{\rm{lnK=\frac{-13.8\times 10^3}{8.314 \times 298}}}$ = -5.569
⇒ lnK = -5.569
2. Taking antilog of both sides, we get: K = e-5.569 = 3.81 × 10⁻³

Solved example 7.32
Calculate K for the reaction of O2 with N2 to give NO at 423 K:
N2 (g) + O2 (g) ⇌ 2NO (g)
∆G⊝ for this reaction is +22.7 kJ/mol
Solution:
1. We have Eq.7.6: $\mathbf\small{\rm{lnK=\frac{-\Delta G^\circleddash}{RT}}}$
• Substituting the known values, we get: $\mathbf\small{\rm{lnK=\frac{-22.7\times 10^3}{8.314 \times 423}}}$ = -6.45
⇒ lnK = -6.45
2. Taking antilog of both sides, we get: K = e-6.45 = 0.00158


Next we will consider the case when the reaction is not at equilibrium. This can be analyzed in 2 steps:
1. When the reaction is not at equilibrium, we cannot put ΔG = 0 in Eq.7.4
    ♦ Also, we cannot put Q = K
• That means, we keep Eq.7.4 as such
2. Consider Eq.7.4: $\mathbf\small{\rm{\Delta G= \Delta G^\circleddash + RT\,lnQ}}$
• The right side has two terms
   ♦ The first term among them is ∆G⊝
         ✰ It will be given in the question
        ✰ Or it can be calculated using Eq.7.5 
   ♦ The second term among them can be calculated using concentrations
• So we can easily calculate ΔG

Solved example 7.33
∆G⊝ for the reaction N2 (g) + 3H2 (g) ⇌ 2NH3 is -32.7 kJ mol-1. Calculate ΔG when the concentrations are as follows:
[N2] =2.00 M, [H2] = 7.00 M, [NH3] = 0.021 M
Temperature = 373 K
Solution:
1. We have Eq.7.4: $\mathbf\small{\rm{\Delta G= \Delta G^\circleddash + RT\,lnQ}}$
The first term on the right = ∆G⊝ = -32.7 kJ mol-1
2. The second term on the right is RT lnQ
• So we have to first find Q
• We have: Q = $\mathbf\small{\rm{\frac{[NH_3]^2}{[N_2][H_2]^3}=\frac{[0.021]^2}{[2.00][7.00]^3}}}$ = 6.428 × 10⁻⁷
• Thus we get: RT lnQ = (8.314 × 373 × ln6.428 × 10-7) = -44214.04 J mol-1 = -44.21 kJ mol-1
3.  Adding the two terms, we get: ΔG = (-32.7 -44.21) = -76.91 kJ mol-1
4. We see that ΔG < 0
So the reaction must proceed spontaneously in the forward direction
5. To confirm this, we have to check the value of K also
• To find K, we can use Eq.7.6: $\mathbf\small{\rm{lnK=\frac{-\Delta G^\circleddash}{RT}}}$
• Substituting the known values, we get: $\mathbf\small{\rm{lnK=\frac{32.7\times 10^3}{8.314 \times 373}}}$ = 10.545
⇒ lnK = 10.545
• Taking antilog of both sides, we get: K = e10.545 = 37987.03
• This value of K is greater than the value of Q that we obtained in (2)
• So the reaction will indeed proceed in the forward direction. Result in (4) is confirmed

Solved example 7.34
Calculate ΔG for the reaction N2(g)+O2(g) ⇌ 2NO(g) under the conditions: T = 423 K, [NO] = 0.0100 M, [O2] = 0.200 M, and [N2] = 1.00 × 10⁻⁴ M. The value of ∆G⊝ for this reaction is +22.7 kJ. In which direction will the reaction proceed to reach equilibrium?
Solution:
1. We have Eq.7.4: $\mathbf\small{\rm{\Delta G= \Delta G^\circleddash + RT\,lnQ}}$
The first term on the right = ∆G⊝ = +22.7 kJ mol-1
2. The second term on the right is RT lnQ
• So we have to first find Q
• We have: Q = $\mathbf\small{\rm{K_c=\frac{[NO]^2}{[N_2][O_2]}=\frac{[0.0100]^2}{[1.00 \times 10^{-4}][0.200]}}}$ = 5
• Thus we get: RT lnQ = (8.314 × 423 × ln5) = 5659.97 J mol-1 = 5.66 kJ mol-1
3.  Adding the two terms, we get: ΔG = (22.7 + 5.66) = +28.36 kJ mol-1
4. We see that ΔG > 0
So the reaction must proceed spontaneously in the backward direction
5. To confirm this, we have to check the value of K also
• To find K, we can use Eq.7.6: $\mathbf\small{\rm{lnK=\frac{-\Delta G^\circleddash}{RT}}}$
• Substituting the known values, we                 get: $\mathbf\small{\rm{lnK=\frac{-22.7\times 10^3}{8.314 \times 423}}}$ = -6.455
⇒ lnK = -6.455
• Taking antilog of both sides, we get: K = e-6.455 = 0.00157
• This value of K is less than the value of Q that we obtained in (2)
• So the reaction will indeed proceed in the backward direction. Result in (4) is confirmed

Solved example 7.35
Calculate a) ∆G⊝ and b) the equilibrium constant for the formation of NO2 from
NO and O2 at 298K
NO (g) + 1⁄2 O2 (g) ⇌ NO2 (g)
where
∆G⊝[NO2] = 52.0 kJ mol-1
∆G⊝[NO] = 87.0 kJ mol-1
∆G⊝[O2] = 0 kJ mol-1
Solution:
Part (a):
1. ∆G⊝ can be obtained by subtracting item (ii) from item (i) below:
(i) Sum of the ∆G⊝ values of all products                                                                   
(ii) Sum of the ∆G⊝ values of all reactants
2. Thus we get:
∆G⊝ = (52.0 - 87) = -35 kJ
Part (b):
1. We have Eq.7.6: $\mathbf\small{\rm{lnK=\frac{-\Delta G^\circleddash}{RT}}}$
• Substituting the known values, we get: $\mathbf\small{\rm{lnK=\frac{35\times 10^3}{8.314 \times 298}}}$ = 14.13
⇒ lnK = 14.13
2. Taking antilog of both sides, we get: K = e14.13 = 1.365 × 10⁶


• In the next section, we will see the factors affecting equilibrium

Previous

Contents

Next

Copyright©2021 Higher secondary chemistry.blogspot.com

Wednesday, March 10, 2021

Chapter 6.15 - Gibbs Free Energy

In the previous section, we saw the second law of thermodynamics. In this section, we will see Gibbs energy

◼ Gibbs energy G is defined as: Eq.6.16: G = H – TS
    ♦ H is the enthalpy of the system
    ♦ T is the temperature of the system
    ♦ S is the entropy of the system
We know that, H, T and S are state functions. So G is also a state function

Now we will see some interesting calculations based on the above equation. It can be written in 11 steps:
1. Initial and final states of the system:
    ♦ At the initial state A, we have: Gsys(A) = Hsys(A) - TSsys(A)
    ♦ At the final state B, we have: Gsys(B) = Hsys(B) - TSsys(B)
2. Now we can find ΔGsys:
ΔGsys = (Gsys(B) - Gsys(A)) = (Hsys(B) - TSsys(B)) - (Hsys(A) - TSsys(A))
⇒ ΔGsys = (Hsys(B) - Hsys(A)) -T(Ssys(B) - Ssys(A))
Thus we get: Eq.6.17: ΔGsys = ΔHsys - T ΔSsys
3. Note that:
    ♦ Eq.6.16 is related to Gibbs energy
    ♦ Eq.6.17 is related to Gibbs energy change
4. Let us do a dimensional analysis of Eq.6.16
• On the right side, we have:
$\mathbf\small{\rm{[Energy]-[Temperature]\times \frac{[Energy]}{[Temperature]}}}$
= [Energy] - [Energy]
= [Energy]
• So we can write: G has the dimensions of energy. In other words, G is a quantity of energy
• ΔG in Eq.6.17 is the difference between two G values. So ΔG is also a quantity of energy
◼ Eq.6.17 is known as the Gibbs equation. It is one of the most important equations in chemistry
5. In earlier sections, we saw that:
    ♦ A decrease in enthalpy (negative ΔH) could be an indication for spontaneity
    ♦ An increase in entropy of the universe is essential for spontaneity
• Now, Eq.6.17 combines both 'change in enthalpy' and 'change in entropy'
6. Let us see how Gibbs energy is related to spontaneity:
• We have the basic equation: Eq.6.14: ΔSuniverse = [ΔSsys + ΔSsurr]
• We calculated the last term Ssurr as follows:
$\mathbf\small{\rm{\Delta S_{surr}=\frac{Heat\,absorbed/released\,by\,surroundings}{T}}}$
7. We saw that:
$\mathbf\small{\rm{Heat\,absorbed/released\,by\,surroundings}}$ is related to the ΔHsys
• The relation can be written in two ways:
(i) If heat ΔHsys is released by the system, that same heat will be absorbed by the surroundings
    ♦ We can write: If ΔHsys is negative, $\mathbf\small{\rm{Heat\,absorbed/released\,by\,surroundings}}$ will be positive but with the same magnitude as ΔHsys
(ii) If heat ΔHsys is absorbed by the system, that same heat will be lost by the surroundings
    ♦ We can write: If ΔHsys is positive, $\mathbf\small{\rm{Heat\,absorbed/released\,by\,surroundings}}$ will be negative but with the same magnitude as ΔHsys
◼ Based on the two points, we can write:
$\mathbf\small{\rm{Heat\,absorbed/released\,by\,surroundings}}$ is the negative of ΔHsys
8. So Eq.6.14 becomes:
ΔSuniverse = [ΔSsys + $\mathbf\small{\rm{\frac{-\Delta H_{sys}}{T}}}$]
• Rearranging this, we get:
TΔSuniverse = TΔSsys - ΔHsys
9. For a spontaneous process, ΔSuniverse will be greater than zero
• T will be always positive because, there are no negative values in the kelvin scale
• Thus we can write:
If the process is spontaneous, (TΔSsys - ΔHsys) will be greater than zero
• (TΔSsys - ΔHsys) can be rearranged as: -(ΔHsys - TΔSsys)
• So we can write:
If the process is spontaneous, -(ΔHsys - TΔSsys) will be greater than zero
• This can be rearranged as:
If the process is spontaneous, (ΔHsys - TΔSsys) will be less than zero
◼ Thus we get an important result:
A process will be spontaneous if: (ΔHsys - TΔSsys) < 0
10. Note that, we are considering the system only. The universe and the surroundings do not come in the equation. So we will drop the subscript 'sys'. We can write:
A process will be spontaneous if: (ΔH - TΔS) < 0
• Also note that, from Eq.6.17, we have: (ΔH - TΔS) = ΔG
◼ So we can write:
A process will be spontaneous if ΔG < 0
11. Consider the equation 6.17: ΔG = ΔH - TΔS
• All three terms are energies
• ΔH is the heat liberated from the system
• But all that ΔH is not available to do work. Because, a quantity of 'TΔS' is being deducted
• The energy remaining after the deduction is the ΔG
• So we can say that, ΔG is the free energy available to do work
◼ For this reason, ΔG is also known as the free energy


Next we will consider the 'possible cases' where a process can be spontaneous or non-spontaneous. It can be written in 6 steps
1. We have Eq.6.17: ΔG = ΔH - TΔS
   ♦ ΔH can be positive or negative
   ♦ ΔS can be positive or negative
   ♦ T can only be positive
        ✰ This is because, there are no negative values in the kelvin scale
2. So four possible cases arise
• Two of them are when ΔH is positive:
   ♦ ΔH positive, ΔS positive
   ♦ ΔH positive, ΔS negative
• Remaining two are when ΔH is negative:
   ♦ ΔH negative, ΔS positive
   ♦ ΔH negative, ΔS negative
3. Let us consider the first case: ΔH +ve, ΔS +ve
• Substituting in Eq.6.17, we get:
ΔG = (+ve) - T(+ve)
• On the right side, first term is positive and second term is negative
   ♦ At small values of T, the first term will be larger
         ✰ Then the result will be a +ve ΔG
   ♦ At large values of T, the first term will be smaller
         ✰ Then the result will be a -ve ΔG
• We know that:
   ♦ -ve ΔG indicates spontaneous process
   ♦ +ve ΔG indicates non-spontaneous process
◼  So for case 1, we can write:
   ♦ ΔH is +ve, ΔS is +ve
         ✰ The process will be spontaneous at large values of T
         ✰ The process will be non-spontaneous at small values of T
◼ This 'case 1' is a special case. It is an endothermic process. That is., heat is absorbed by the system. We see that, if T is increased to a high level, even an endothermic process can be made to take place spontaneously
4. Let us consider the second case: ΔH +ve, ΔS -ve
• Substituting in Eq.6.17, we get:
ΔG = (+ve) - T(-ve)
• On the right side, first term is positive and second term is also positive
   ♦ So whatever be the value of T, ΔG will be always +ve
• We know that:
   ♦ -ve ΔG indicates spontaneous process
   ♦ +ve ΔG indicates non-spontaneous process
◼  So for case 2, we can write:
   ♦ ΔH is +ve, ΔS is +ve
         ✰ Whatever be the value of T, the process will be always non-spontaneous
5. Let us consider the third case: ΔH -ve, ΔS +ve
• Substituting in Eq.6.17, we get:
ΔG = (-ve) - T(+ve)
• On the right side, first term is negative and second term is also negative
   ♦ So whatever be the value of T, ΔG will be always -ve
• We know that:
   ♦ -ve ΔG indicates spontaneous process
   ♦ +ve ΔG indicates non-spontaneous process
◼  So for case 3, we can write:
   ♦ ΔH is -ve, ΔS is +ve
         ✰ Whatever be the value of T, the process will be always spontaneous
6. Let us consider the fourth case: ΔH -ve, ΔS -ve
• Substituting in Eq.6.17, we get:
ΔG = (-ve) - T(-ve)
• On the right side, first term is negative and second term is positive
   ♦ At small values of T, the first term will be larger
         ✰ Then the result will be a -ve ΔG
   ♦ At large values of T, the first term will be smaller
         ✰ Then the result will be a +ve ΔG
• We know that:
   ♦ -ve ΔG indicates spontaneous process
   ♦ +ve ΔG indicates non-spontaneous process
◼  So for case 4, we can write:
   ♦ ΔH is -ve, ΔS is -ve
         ✰ The process will be spontaneous at small values of T
         ✰ The process will be non-spontaneous at large values of T
◼ This 'case 4' is a special case. It is an exothermic process. That is., heat is released by the system. We see that, if T is increased to a high level, even an exothermic process can be made to take place non-spontaneously


Let us see some solved examples:
Solved example 6.30
A reaction, A + B → C + D + q is found to have a positive entropy change. The
reaction will be
(i) possible at high temperature
(ii) possible only at low temperature
(iii) not possible at any temperature
(v) possible at any temperature
Solution:
1. The given equation is: A + B → C + D + q
• It is not a thermochemical equation because, the energy is written along with the equation
• The products constitute of: C, D and q
    ♦ That means, q is also produced. So it is an exothermic reaction
• We can write the thermochemical equation as: A + B → C + D; ΔH = -q
2. So ΔH is negative. We are given that, ΔS is positive
• Thus this process falls under case 3: ΔH -ve, ΔS +ve
3. We can write:
Whatever be the temperature, the given process will be always spontaneous

Solved example 6.31
For the reaction at 298 K,
2A + B → C
∆H = 400 kJ mol-1 and ∆S = 0.2 kJ K-1 mol-1
At what temperature will the reaction become spontaneous considering ∆H and ∆S to be constant over the temperature range
Solution:
1. We have E.6.17: ΔG = ΔH - TΔS
• For the reaction to be spontaneous, ∆G must be less than zero
2. To find the temperature at which the reaction just becomes spontaneous, we put: ∆G = 0
• Thus we get: 0 = ΔH - TΔS
3. Substituting the known values, we get:
0 = 400 (kJ mol-1) - [T (K) × 0.2 (kJ K-1 mol-1)]
⇒ T = 2000 K


Solved example 6.32
For the reaction,
2Cl(g) → Cl2(g), what are the signs of ∆H and ∆S ?
Solution:
1. Sign of ∆H:
• In this process, individual Cl atoms combine to form Cl2 molecules
    ♦ Bond enthalpy will be released
    ♦ So ∆H will be negative
2. Sign of ∆S:
• In the initial state, there are more number of particles
• In the final state, number of particles become half
    ♦ This reduces the disorder/randomness
    ♦ So S decreases
    ♦ Thus ∆S will be negative 

Solved example 6.33
For the reaction
2A(g) + B(g) → 2D(g)
∆U = –10.5 kJ and ∆S = –44.1 J K-1
Calculate ∆G for the reaction, and predict whether the reaction may occur
spontaneously
Solution:
• We have seen this type of problems in section 6.3
• We will solve this problem using Method I only
• The reader may try Method II also
Method I:
1. The given balanced equation is:
2A(g) + B(g) → 2D(g)
• Let A be the initial state
• In this state, the system is:
    ♦ 2 mol of A and 1 mol B at 298 K and P atm pressure
2. Let B be the final state
• In this state, the system is:
    ♦ 2 mol of D at 298 K and P atm pressure
3. We have seen that:
    ♦ Enthalpy in the initial state, HA = (UA + PVA)
    ♦ Enthalpy in the final state, HB = (UB + PVB)
    ♦ So enthalpy change = (HB - HA) = [(UB - UA) + P(VB - VA)]
4. Here, (UB - UA) is given as -10.5 KJ
• So we can write:
[(-10.5) + P(VB - VA)] = HB - HA
5. In the above equation, there are 3 terms: (-10.5), P(VB - VA) and (HB - HA)
• So, if we can find P(VB - VA), we can calculate (HB - HA)
6. P(VB - VA) can be calculated in steps:
(i) From ideal gas equation, we have:
    ♦ PAVA = nARTA
    ♦ PBVB = nBRTB
• In our present case:
    ♦ PA = PB = P
    ♦ TA = TB = T = 298 K
(ii) So P(VB - VA) = (nB - nA)RT
    ♦ nA = number of gaseous moles in the initial state = total number of gaseous moles of A and B = (2+1) = 3
    ♦ nB = number of gaseous moles in the final state = number of gaseous moles of D = 2
    ♦ So (nB - nA) = (2 - 3) = -1
(iii) Substituting the known values, the right side of (ii) becomes:
(-1 mol) × (8.3 J mol-1 K-1) × (298 K) = -2473.4 J = -2.473 kJ
7. We can use the value obtained above, in the place of P(VB - VA) in (4)
• We get: [(-10.5) -2.473] = (HB - HA)
• So (HB - HA) = ΔH = -[12.973] kJ
8. Now we use Eq.6.17: ΔG = ΔH - TΔS
• Substituting the known values, we get:
ΔG = -12.973 × 10³ - (298  × -44.1) = 168.8 J
• Since this value is positive, the reaction is not spontaneous

Solved example 6.34
Calculate the entropy change in surroundings when 1.00 mol of H2O(l) is formed
under standard conditions. ΔH⊖f  = –286 kJ mol-1
Solution:
1. Given that, ΔH⊖f  = –286 kJ mol-1
• So 286 kJ will be absorbed by the surroundings
2. We have: $\mathbf\small{\rm{\Delta S_{surr}=\frac{Heat\,absorbed\,by\,surroundings}{T}}}$
• The 286 should be given a +ve sign because, heat content of the surroundings increases due to the absorption
• We get: $\mathbf\small{\rm{\Delta S_{surr}=\frac{286 \times 10^3}{298}}}$ = 959.73 J K-1


• In the next chapter, we will see equilibrium


Previous

Contents

Next

Copyright©2021 Higher secondary chemistry.blogspot.com
 

Monday, March 8, 2021

Chapter 6.14 - Second Law of Thermodynamics

In the previous section, we saw some basics about entropy. In this section, we will see the second law of thermodynamics

◼ The second law of thermodynamics states that:
When a spontaneous process takes place, the entropy of the universe always increases
• This can be explained in 6 steps:
1. Universe is equivalent to (system + surroundings)
• So entropy of universe, Suniverse = Ssys + Ssurr
2. Initial and final states:
    ♦ At the initial state A, we have: Suniverse(A) = Ssys(A) + Ssurr(A)
    ♦ At the final state B, we have: Suniverse(B) = Ssys(B) + Ssurr(B)
3. The second law states that, the entropy of the universe always increases
    ♦ That means: Suniverse(B) must be larger than Suniverse(A)
    ♦ That means: [Suniverse(B) - Suniverse(A)] must be greater than zero
    ♦ That means: ΔSuniverse must be greater than zero
4. We have:
ΔSuniverse = [Suniverse(B) - Suniverse(A)]
⇒ ΔSuniverse = [(Ssys(B) + Ssurr(B)) - (Ssys(A) + Ssurr(A))]
⇒ ΔSuniverse = [(Ssys(B) - Ssys(A)) + (Ssurr(B) - Ssurr(A))]
• Thus we get Eq.6.14: ΔSuniverse = [ΔSsys + ΔSsurr]
5. So based on the second law, we can write:
For a process to be spontaneous, ΔSuniverse > 0
6. From (4), it is clear that:
To calculate ΔSuniverse, we must know ΔSsys and ΔSsys
    ♦ We know how to find ΔSsys
        ✰ see solved examples 6.23 and 6.24 of the previous section
    ♦ Later in this section, we will see the method to find ΔSsurr


• Using the second law, we can predict the direction of a process
• For that, we look at the entropy in two directions
1. First we consider the direction from left to right
• When the process proceeds in this direction, if we get ΔSuniverse > 0, then the  process will occur spontaneously in this direction
• That means, no external energy is required for the reaction to proceed from left to right
• The opposite is also true:
When the process proceeds in this direction, if we get ΔSuniverse < 0, then the  process will not occur spontaneously in this direction
• That means, external energy is required for the reaction to proceed from left to right
2. Next we consider the direction from right to left
• When the process proceeds in this direction, if we get ΔSuniverse > 0, then the  process will occur spontaneously in this direction
• That means, no external energy is required for the reaction to proceed from right to left 
• The opposite is also true:
When the process proceeds in this direction, if we get ΔSuniverse < 0, then the process will not occur spontaneously in this direction
• That means, external energy is required for the reaction to proceed from right to left


• As mentioned in (6) above, our next aim is to find ΔSsurr. It can be written in 5 steps:
1. Initial and final entropies:
• We have:
    ♦ Entropy of the surroundings in the initial state = Ssurr(A)
    ♦ Entropy of the surroundings in the final state = Ssurr(B)
2. Then change in entropy ΔSsurr = Ssurr(B) – Ssurr(A)
3. This change in entropy is due to the absorption of a heat Q
• This Q can be large or small:
• If Q is large, Ssurr(B) will be large
    ♦ (Because, large Q causes large disorder)
    ♦ Then ΔSsurr will be large
• If Q is small, Ssurr(B) will be small
    ♦ Then ΔSsurr will be small
◼ We can write:
ΔSsurr is directly proportional to Q
4. Based on experiments, scientists obtained another information. It can be written in 6 steps:
(i) Let the surroundings be at a lower temperature T1
    ♦ Let a heat Q be added to the surroundings
    ♦ Let the resulting change in entropy be ΔSsurr(1)
(ii) Let the surroundings be at a higher temperature T2
    ♦ Let the same heat Q be added to the surroundings
    ♦ Let the resulting change in entropy be ΔSsurr(2)
(iii) Here we see an interesting fact:
    ♦ ΔSsurr(2) will be smaller than ΔSsurr(1)
• Let us analyze the reason:
(iv) At the lower temperature T1, the surroundings is somewhat calm
    ♦ To the calm surroundings, we are adding Q
• At the higher temperature T2, the surroundings will already have some disorder
    ♦ To that disordered surroundings, we are adding the same Q
(v) Observing the 'change in disorder' at two temperatures:
    ♦ The 'change in disorder' created by Q at the lower temperature T1
    ♦ Will be more observable than
    ♦ The 'change in disorder' created by the same Q at the higher temperature T2
• This is because, at T2, the surroundings already have some disorder
(vi) So we can write:
    ♦ ΔSsurr(1) at lower temperature is high
    ♦ ΔSsurr(2) at higher temperature is low
• That means:
ΔSsurr is inversely proportional to the temperature
5. So we have two information:
    ♦ From (3), we have: ΔSsurr is directly proportional to Q
    ♦ From (4), we have: ΔSsurr is inversely proportional to T
◼ Combining the two, we get:
Eq.6.15: $\mathbf\small{\rm{\Delta S_{surr}=\frac{Q}{T}}}$
• From this equation, it is clear that, units of entropy is J K-1


Now that we know the two terms of Eq.6.14, we will see a practical application of the second law. The following five solved example demonstrates the application

Solved example 6.25
Prove that:
   ♦ Ice will not melt spontaneously at -5 ०C
   ♦ Ice will begin to melt spontaneously at 0 ०C
   ♦ Ice will melt spontaneously at 5 ०C
Given that: For the process H2O(s)  → H2O(l), ΔS⊖ = 22.0 J K-1 mol-1
Solution:
1. We have to analyze the process: H2O(s)  → H2O(l)
• We have seen this process in an earlier section. (see 'Enthalpy changes during phase transformations' in section 6.6)
• The thermochemical equation is: H2O(s)  → H2O(l) ΔH⊖fusion = 6.00 kJ mol-1
2. To determine whether a process is spontaneous, we have to apply Eq.6.14:
ΔSuniverse = [ΔSsys + ΔSsurr]
3. First we will find ΔSsys
• But it is already given in the question. ΔSsys = ΔS⊖ = 22.0 J K-1 mol-1
• Let us consider one mol ice. Then we can ignore the 'mol-1'
• We can write:
For our present system, ΔSsys = 22.0 J K-1
4. Next we will find ΔSsurr
• We have Eq.6.15: $\mathbf\small{\rm{\Delta S_{surr}=\frac{Q}{T}}}$
5. From the thermochemical equation, we see that, when 1 mol ice melts, the surroundings will lose 6.00 kJ energy
   ♦ Let QA be the initial heat of the surroundings
   ♦ Let QB be the final heat of the surrounding
• QB will be less than QA because, heat is lost by the surroundings
   ♦ This lost heat is used up for melting the ice
• The change of heat = (QB -QA) = -6 kJ
   ♦ The -ve sign is to be given for 6 because, QB will be less than QA
6. Substituting the known values in (4), we get:
$\mathbf\small{\rm{\Delta S_{surr}=\frac{-6000(J)}{T(K)}}}$
7. Now the result in (2) becomes:
ΔSuniverse = [22.0 J K-1 + $\mathbf\small{\rm{\frac{-6000(J)}{T(K)}}}$]
• Now we can take up each case
8. Case 1: When temperature is -5 ०C
• -5 ०C is 268 K
• So the result in (7) becomes:
ΔSuniverse = [22.0 J K-1 + $\mathbf\small{\rm{\frac{-6000(J)}{268(K)}}}$] = -0.3881
◼  This is a negative value. So we can write:
If this process takes place at -5 ०C, entropy of the universe decreases
• Any process which causes a decrease in entropy of the universe will not be spontaneous
   ♦ So the process H2O(s)  → H2O(l) is not spontaneous at -5 ०C
   ♦ The reverse process H2O(l)  → H2O(s) will be spontaneous at -5 ०C
9. Case 2: When temperature is 0 ०C
• 0 ०C is 273 K
• So the result in (7) becomes:
ΔSuniverse = [22.0 J K-1 + $\mathbf\small{\rm{\frac{-6000(J)}{273(K)}}}$] = 0.022
◼  This value is close to zero. So we can write:
If this process takes place at 0 ०C, entropy of the universe neither increases nor decreases
• Any process which neither increases nor decreases the entropy of the universe will be at equilibrium
   ♦ So the process H2O(s)  → H2O(l) is at equilibrium at 0 ०C
   ♦ If the temperature rises just above 0 ०C, melting will begin
10. Case 3: When temperature is +5 ०C
• +5 ०C is 278 K
• So the result in (7) becomes:
ΔSuniverse = [22.0 J K-1 + $\mathbf\small{\rm{\frac{-6000(J)}{278(K)}}}$] = 0.4172 J K-1
◼  This is a positive value. So we can write:
If this process takes place at +5 ०C, entropy of the universe increases
• Any process which causes an increase in entropy of the universe will be spontaneous
   ♦ So the process H2O(s)  → H2O(l) is spontaneous at +5 ०C
   ♦ The reverse process H2O(l)  → H2O(s) will not be spontaneous at +5 ०C

Solved example 6.26
The process White Tin(s) → Gray Tin(s) occurs when the surrounding temperature falls just below 13.2 ०C. The ΔH for the process is -2.1 kJ mol-1. What is the ΔS for the process? Which one has greater order? White tin or Gray tin?
Solution:
1. We have to analyze the process: White Tin(s)  → Gray Tin(s); ΔH = -2.1 kJ mol-1
2. To determine whether a process is spontaneous, we have to apply Eq.6.14:
ΔSuniverse = [ΔSsys + ΔSsurr]
3. First we will write ΔSsys
• We have: ΔSsys = S[Product] - S[Reactants]
⇒ ΔSsys = S[Gray] - S[White]
4. Next we will find ΔSsurr
We have Eq.6.15: $\mathbf\small{\rm{\Delta S_{surr}=\frac{Q}{T}}}$
5. From the thermochemical equation, we see that, when one mol white Tin gets converted into one mol gray Tin, 2.1 kJ is released
• So the surroundings will gain 2.1 kJ energy
   ♦ Let QA be the initial heat of the surroundings
   ♦ Let QB be the final heat of the surrounding
• QB will be greater than QA because, heat is gained by the surroundings
• The change of heat = (QB - QA) = +2.1 kJ
   ♦ The +ve sign is to be given for 2.1 because, QB will be greater than QA
6. Substituting the known values in (4), we get:
$\mathbf\small{\rm{\Delta S_{surr}=\frac{+2100(J)}{T(K)}}}$
7. Now the result in (2) becomes:
ΔSuniverse = [S[Gray] - S[White] + $\mathbf\small{\rm{\frac{+2100(J)}{T(K)}}}$]
8. Given that, the conversion just begins at 13.2 ०C
    ♦ That means, at 13.2 ०C, the system is in equilibrium
    ♦ That means, at 13.2 ०C, there is no change in entropy of the universe
    ♦ That means, at 13.2 ०C, ΔSuniverse = 0
• 13.2 ०C is equal to (273.15 + 13.2) = 286.4 K
9. So the result in (7) becomes:
0 = [S[Gray] - S[White] + $\mathbf\small{\rm{\frac{+2100(J)}{286.4(K)}}}$]
⇒ S[Gray] - S[White] = $\mathbf\small{\rm{\frac{-2100(J)}{286.4(K)}}}$ = -7.33 J K-1
10. So we can write:
ΔS for the process White Tin(s) → Gray Tin(s) is -7.33 J K-1
11. The -ve value of the ΔS indicates that, the entropy decreases
    ♦ That means, the reactant has greater entropy
    ♦ That means, the reactant has greater disorder
    ♦ That means the product has greater order
• So we can write:
Gray Tin has greater order

Solved example 6.27
The process Orthorhombic Sulfur(s) → Monoclinic Sulfur(s) occurs when the surrounding temperature rises just above 95.3 ०C. The ΔH for the process is +0.401 kJ mol-1. What is the ΔS for the process? Which one has greater order? Orthorhombic Sulfur or Monoclinic Sulfur?
Solution:
1. We have to analyze the process:
Orthorhombic Sulfur(s) → Monoclinic Sulfur(s); ΔH = +0.401 kJ mol-1
2. To determine whether a process is spontaneous, we have to apply Eq.6.14:
ΔSuniverse = [ΔSsys + ΔSsurr]
3. First we will write ΔSsys
• We have: ΔSsys = S[Product] - S[Reactants]
⇒ ΔSsys = S[Mono] - S[Ortho]
4. Next we will find ΔSsurr
• We have Eq.6.15: $\mathbf\small{\rm{\Delta S_{surr}=\frac{Q}{T}}}$
5. From the thermochemical equation, we see that, when one mol ortho gets converted into one mol mono, 0.401 kJ is absorbed
• So the surroundings will lose 0.401 kJ energy
   ♦ Let QA be the initial heat of the surroundings
   ♦ Let QB be the final heat of the surrounding
• QB will be lesser than QA because, heat is lost by the surroundings
• The change of heat = (QB - QA) = -0.401 kJ
   ♦ The -ve sign is to be given for 0.401 because, QB will be lesser than QA
6. Substituting the known values in (4), we get:
$\mathbf\small{\rm{\Delta S_{surr}=\frac{-401(J)}{T(K)}}}$
7. Now the result in (2) becomes:
ΔSuniverse = [S[Mono] - S[Ortho] + $\mathbf\small{\rm{\frac{-401(J)}{T(K)}}}$]
8. Given that, the conversion just begins at 95.3 ०C
    ♦ That means, at 95.3 ०C, the system is in equilibrium
    ♦ That means, at 95.3 ०C, there is no change in entropy of the universe
    ♦ That means, at 95.3 ०C, ΔSuniverse = 0
• 95.3 ०C is equal to (273.15 + 95.3) = 368.45 K
9. So the result in (7) becomes:
0 = [S[Mono] - S[Ortho] + $\mathbf\small{\rm{\frac{-401(J)}{368.45(K)}}}$]
⇒ S[Mono] - S[Ortho] = $\mathbf\small{\rm{\frac{401(J)}{368.45(K)}}}$ = 1.09 J K-1
10. So we can write:
ΔS for the process Orthorhombic Sulfur(s) → Monoclinic Sulfur(s) is 1.09 J K-1
11. The +ve value of the ΔS indicates that, the entropy increases
    ♦ That means, the product has greater entropy
    ♦ That means, the product has greater disorder
    ♦ That means the reactant has greater order
• So we can write:
Orthorhombic sulfur has greater order

Solved example 6.28
Calculate the entropy change for oxidation of iron, 4Fe(s) + 3O2 (g) → 2Fe2O3(s). Inspite of negative entropy change of this reaction, why is the reaction spontaneous at 298 K?
(ΔH⊖r of the reaction is –1648 × 103 J mol-1)
Solution:
1. We have already calculated the entropy change for this reaction
(see solved example 6.23 of the previous section)
• We got: ΔS⊖r = -549.74 J K-1
2. To determine whether a process is spontaneous, we have to apply Eq.6.14:
ΔSuniverse = [ΔSsys + ΔSsurr]
3. The result in (1) is ΔSsys
So we have: ΔSsys = -549.74 J K-1
4. Next we will find ΔSsurr
• We have Eq.6.15: $\mathbf\small{\rm{\Delta S_{surr}=\frac{Q}{T}}}$
5. From the data given, we see that, during the reaction, 1648 × 103 J is released
• So the surroundings will gain 1648 × 103 J energy
   ♦ Let QA be the initial heat of the surroundings
   ♦ Let QB be the final heat of the surrounding
• QB will be greater than QA because, heat is gained by the surroundings
• The change of heat = (QB - QA) = +1648 × 103 J
   ♦ The +ve sign is to be given for 1648 × 103 because, QB will be greater than QA
6. Substituting the known values in (4), we get:
$\mathbf\small{\rm{\Delta S_{surr}=\frac{+1648 × 10^3 (J)}{T(K)}}}$
7. Now the result in (2) becomes:
ΔSuniverse = [-549.74 J K-1 + $\mathbf\small{\rm{\frac{+1648 × 10^3(J)}{T(K)}}}$]
8. We have to examine the reaction at 298 K. So T = 298 K
9. So the result in (7) becomes:
ΔSuniverse = [-549.74 J K-1 + $\mathbf\small{\rm{\frac{+1648 × 10^3(J)}{298(K)}}}$]
⇒ ΔSuniverse = 4980.5
10. This is a positive value
• That means, the entropy of the universe increases
• So the process will be spontaneous

Solved example 6.29
Calculate the temperature at which the following reaction becomes spontaneous:
CaCO3(s) → CaO(s) + CO2(g); ΔH⊖r = +179 kJ mol-1
Solution:
1. We have to analyze the process:
CaCO3(s) → CaO(s) + CO2(g); ΔH⊖r = +179 kJ mol-1
2. To determine whether a process is spontaneous, we have to apply Eq.6.14:
ΔSuniverse = [ΔSsys + ΔSsurr]
3. We have: ΔSsys = S[Product] - S[Reactants]
⇒ ΔS⊖r = S⊖[CaO(s)] + S⊖[CO2(g)] - S⊖[CaCO3(s)]
⇒ ΔS⊖r = 39.75 + 213.74 - 92.9 = 160.59
4. Next we will find ΔSsurr
• We have Eq.6.15: $\mathbf\small{\rm{\Delta S_{surr}=\frac{Q}{T}}}$
5. From the data given, we see that, during the reaction, 179 × 103 J is absorbed
• So the surroundings will lose 179 × 103 J energy
   ♦ Let QA be the initial heat of the surroundings
   ♦ Let QB be the final heat of the surrounding
• QB will be lesser than QA because, heat is lost by the surroundings
• The change of heat = (QB - QA) = -179 × 103 J
   ♦ The -ve sign is to be given for 179 × 103 because, QB will be lesser than QA
6. Substituting the known values in (4), we get:
$\mathbf\small{\rm{\Delta S_{surr}=\frac{-179 × 10^3 (J)}{T(K)}}}$
7. Now the result in (2) becomes:
ΔSuniverse = [160.59 J K-1 + $\mathbf\small{\rm{\frac{-179 × 10^3(J)}{T(K)}}}$]
8. We want ΔSuniverse to be positive. Only then will the reaction become spontaneous
• The sign of ΔSuniverse is decided by two terms:
   ♦ 160.59 and $\mathbf\small{\rm{\frac{-179 × 10^3(J)}{T(K)}}}$
• 179 × 103 is very large when compared to 160.59
   ♦ So we will want a large 'T' to make the second term small
9. We will first find the equilibrium temperature by equating ΔSuniverse to zero. We get:
0 = [160.59 J K-1 + $\mathbf\small{\rm{\frac{-179 × 10^3(J)}{T(K)}}}$]
⇒ T = 1114.64 K
10. If we raise T a little above 1114.64 K, the reaction will become spontaneous


• In the next section, we will see Gibbs energy


Previous

Contents

Next

Copyright©2021 Higher secondary chemistry.blogspot.com


Thursday, March 4, 2021

Chapter 6.12 - Spontaneity

In the previous section, we saw details about endothermic or exothermic nature of enthalpy of solution. In this section, we will see spontaneity

• Basics of spontaneity can be written in 15 steps:
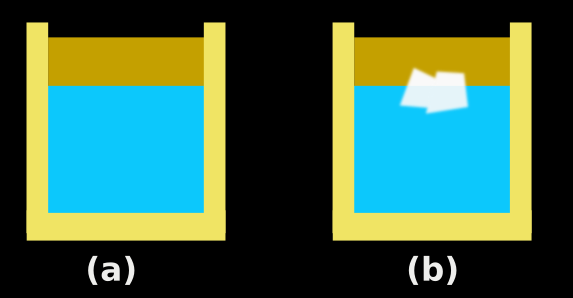
1. Consider the open vessel in fig.6.20(a) below. It contains some water at room temperature 25 ०C

Second law of thermodynamics
Fig.6.20

2. We know that, a mass of water is nothing but a collection of H2O molecules
• All those molecules are in random motion
• So all those molecules have kinetic energies
• The temperature of the water mass is the average kinetic energy of the molecules
3. Due to the random motion, the molecules are constantly colliding with each other
• Imagine that, due to the collisions, some molecules near the surface of the water, looses considerable kinetic energies
    ♦ Then the temperature of those molecules will fall
• Let the temperature fall to such a low level that, those molecules freeze and become ice
4. Remember that, loss of kinetic energy is due to collisions. The kinetic energies lost by the molecules will be gained by the surrounding molecules
• So, when the molecules freeze and become ice, the surrounding molecules become warm
• We will get some ice at the center of the vessel and that ice will be surrounded by warm water. This is shown in fig.6.20(b)
5. Formation of ice in this manner, does not defy the law of conservation of energy because,
    ♦ there is no destruction of energy
    ♦ also, no energy is created
• The energy lost by some molecules is gained by some other molecules
6. Even though the law of conservation is obeyed, we never see such a spontaneous formation of ice at room temperature


• A Spontaneous process is the one which do not require external source of energy to proceed
• For a process to be called spontaneous, it is not necessary that it occurs at a high speed
• A slow process can also be called spontaneous if it does not require external supply of energy
• The reaction between hydrogen and oxygen is an example of a slow spontaneous process
    ♦ A mixture of H2 and O2 can be left undisturbed in a container for many years
    ♦ There will not be any noticeable effects
    ♦ But the reaction will be taking place all the time, with out any aid of external energy

7. The spontaneous formation of ice in this manner requires heat to flow (without external help) from a cold object to a hot object
◼ We never observe such a flow. What we observe is the flow of heat from hot object to cold object
• We can say:
    ♦ The spontaneous process proceeds only in one direction: Ice → Water
    ♦ We never see the spontaneous process: Water → Ice
        ✰ For this process, we have to supply energy through a refrigerator
◼ In fact, all naturally occurring processes (physical or chemical) will tend to proceed in one direction only
8. Let us see another example:
• If a canister containing some gas is opened, the gas molecules will spontaneously spread out into the whole volume of the room
• We never see the gas molecules in the room to enter back spontaneously into the canister
9. One more example:
• Consider the burning of carbon
• During the process, carbon combines with oxygen to give carbon dioxide
    ♦ That is: C + O2 → CO2
    ♦ This is a spontaneous process
    ♦ Once the carbon is ignited, no external energy is required to keep the process going
• In the reverse process, carbon and oxygen is obtained from carbon dioxide
    ♦ That is: CO2 → C + O2
    ♦ This reverse process is not spontaneous
    ♦ Energy is required to accomplish this reverse process
10. We see that, all spontaneous processes proceed in one direction only
• We want to answer this question:
Why all spontaneous processes proceed only in one direction?
11. To find the answer, we consider some common phenomena that we see in our day to day life
(i) Flowing of water
• The flow of water starts from top of hill and ends at the ground level
• During the flow, the potential energy stored in the water is continuously released in the form of kinetic energy
• When the water reaches the ground level, it’s potential energy will be zero. This is because, all the potential energy is released (in the form of kinetic energy) into the surroundings
(ii) Stone falling from a height
• The fall of stone starts from a higher level and ends at the ground level
• During the fall, the potential energy stored in the stone is continuously released in the form of kinetic energy
• When the stone reaches the ground level, it’s potential energy will be zero. This is because, all the potential energy is released (in the form of kinetic energy) into the surroundings
12. The flow of water and fall of stone are spontaneous processes. They do not require any aid of external energies
• So we are inclined to think that:
All processes in which there is a ‘release of stored energy’ will be spontaneous
13. We know that, in exothermic reactions, the stored chemical energy is released as heat energy
• So we are inclined to think that:
    ♦ All exothermic reactions are spontaneous
    ♦ The reverse of an exothermic reaction is endothermic
        ✰ It involves absorbtion of energy
    ♦ So a spontaneous reaction proceeds in the exothermic direction only
• Let us examine whether this is true
14. Some thermochemical equations are given below:
(i) 1/2N2(g) + 3/2H2(g) → NH3(g); ΔH⊖r = – 46.1 kJ mol-1
(ii) 1/2H2(g) + 1/2Cl2(g) → HCl(g); ΔH⊖r = – 92.32 kJ mol-1
(iii) H2 + 1/2O2(g) → H2O(l); ΔH⊖r = –285.8 kJ mol-1
• The above three reactions are spontaneous
• The negative sign of ΔH⊖r shows that, they are exothermic reactions
◼ So we become even more inclined to think that:
    ♦ All exothermic reactions are spontaneous
    ♦ Being exothermic is the only criterion for spontaneity
15. But before making a decision, let us see two more thermochemical equations:
(i) 1/2N2(g) + O2(g) → NO2 (g); ΔH⊖r = +33.2 kJ mol-1
(ii) C(graphite, s) + 2S(l) → CS2(l); ΔH⊖r = +128.5 kJ mol-1
• The above two reactions are spontaneous
• The positive sign of ΔH⊖r shows that, they are endothermic reactions


◼ So it is impossible to conclude that:
Being exothermic is the only criterion for a reaction to be spontaneous
• Scientists became convinced that, there are some other factors also playing major roles
• Researches in this direction lead to the discovery of entropy and the second law of thermodynamics
• We will see them in the next section


Previous

Contents

Next

Copyright©2021 Higher secondary chemistry.blogspot.com